

📍Biophysics PhD Candidate @PolizziLab 🎓 B.A. Biophysics @JohnsHopkins '22. On BlueSky until further notice 🦋
Cambridge, MA
Joined October 2014
- Tweets114
- Following552
- Followers504
- Likes2.9K




Ben Fry retweeted

I have postdoc positions available to lead two new exciting projects! One is on in situ protein sequencing (collab with Ed Boyden at MIT) and the other is on modeling cryptic pockets for drug discovery (collab with Scripps and MGH). Reach out if interested! Details below
Ben Fry retweeted

Excited to share a preprint of my main PhD work in @nickpolizzi_'s lab - introducing LUCI-tag, a new orthogonal self-labeling protein tag based on K-Ras(G12C) and its covalent inhibitors! biorxiv.org/content/10.64898… (1/16)

Ben Fry retweeted
Do you use self-labeling proteins (SLPs) like HaloTag and SNAP-tag? Then check out Jody's new preprint to learn about LUCI-tag! Jody built LUCI-tag starting from K-Ras(G12C), using protein design to erase K-Ras from biology 🤯!

Excited to share a preprint of my main PhD work in @nickpolizzi_'s lab - introducing LUCI-tag, a new orthogonal self-labeling protein tag based on K-Ras(G12C) and its covalent inhibitors! biorxiv.org/content/10.64898… (1/16)
In case you missed it, I presented our recent Nature publication at the @ml4proteins seminar series!
The recording is now available on YouTube:
youtube.com/watch?v=BdILCc45…
Link to the paper:
nature.com/articles/s41586-0…
Ben Fry retweeted

We’re excited to host @benf549
tomorrow at 4-5 PM ET, for a presentation on his recent work on Nature
nature.com/articles/s41586-0…

Ben Fry retweeted

Today in @Nature we are excited to share the final version of Dyna-1 and *all* of the datasets we curated! New to the manuscript is prospective experimental validation. Congrats to @HWaymentSteele @DorotheeKern @sokrypton and team!!!
🔗nature.com/articles/s41586-0…
📑rdcu.be/fy2Ax

Ben Fry retweeted

De novo design of small-molecule–induced conformational change
1. Chang & Polizzi present a general, de novo strategy to turn a static small-molecule binder into a ligand-induced conformational switch by adding a mobile “lid” domain that closes behind the ligand, inspired by natural lid mechanisms (e.g., streptavidin, Abl kinase, GPCR loops).
2. Starting scaffold: EPIC, a hyperstable 17 kDa de novo exatecan binder (Tm > 95 °C, Kd ~1.2 nM). Key design opportunity: ~50 Å2 of exatecan remains solvent-exposed in EPIC, including a charged amine that can be “captured” by a designed lid.
3. Core design concept: preserve preorganization in the original binding core (fast association) while introducing induced-fit behavior through a tethered lid. The lid must (i) remain open enough for ligand entry (negative design) and (ii) become stabilized in a closed register only when ligand is present.
4. Computational pipeline: QBITS (query-based interfacial tertiary structures) mines the PDB for recurring helical interface geometries matching a two-helix query near the ligand-exposed region; RFdiffusion extends candidate lid placements; a flexible (GGS)n linker (n=4–7) tethers lid to binder; LASErMPNN designs lid and lid–binder interface residues.
5. Key screening innovation: AlphaFold3 is used in a paired bound/unbound ranking scheme. Designs are prioritized where AF3 predicts confident lid placement only in the ligand-bound state (low interfacial predicted aligned error, iPAE) but is uncertain in the apo state (high iPAE), using ΔiPAE as a proxy for ligand-induced closure.
6. Experimental throughput: only 24 designs were built and tested; all expressed well in E. coli and retained exatecan binding (Kd < 1 µM). Lid closure was detected via (a) fluorescein-on-lid anisotropy shifts and (b) ligand-dependent protection from tryptic cleavage (limited proteolysis as a sensitive closure readout).
7. Lead designs d6.4 and d6.9 achieved sub-nanomolar affinity via lid closure: Kd ~200 pM (d6.4) and ~510 pM (d6.9). The lid adds favorable lid–ligand energetics (estimated ΔΔG contributions of ~−3.2 and −2.3 kcal/mol) and fully buries exatecan (0 Å2 exposed SASA), enabling recognition of essentially the entire ligand surface.
8. Selectivity jump: compared to EPIC, lidded designs strongly discriminate exatecan from the close analog belotecan. EPIC binds belotecan at ~290 nM, while lidded proteins bind belotecan very weakly (Kd > 10 µM), yielding >1:20,000 specificity and >100-fold improvement in specificity ratio vs the parent binder.
9. Structural validation: crystal structures of d6.4:exatecan and d6.9:exatecan closely match AF3 predictions (Cα RMSD ~0.3–0.4 Å). Designed features are observed, including a resolved salt bridge between exatecan’s amine and a designed Glu (Glu41) that helps lock lid register upon binding.
10. Kinetics engineering + biosensing: lid occlusion yields tunable kon/koff and very long residence times (inferred koff down to ~2×10^-7 s^-1; ~months for d6.4). Kinetics are rationally tuned by (i) a single interface mutation (M93E) that opens the apo ensemble and speeds kon ~30-fold while retaining very tight binding, and (ii) lid truncations (via circular permutation) that systematically increase kon. A circularly permuted/truncated variant (d6.4cpΔ11) was converted into a genetically encodable FRET sensor (mTurquoise2–binder–mVenus) with low-nanomolar apparent Kd (~1.5 nM) and robust function in saliva, urine, and serum.
💻Code: github.com/npolizzi/Qbits/re…
📜Paper: biorxiv.org/content/10.64898…
#ProteinDesign #DeNovoProteinDesign #ComputationalBiology #StructuralBiology #Biosensors #DrugDiscovery #AlphaFold3 #RFDiffusion #FRET #Kinetics

LASErMPNN used to search sequence space for dynamic lid sequences!

Ben Fry retweeted
The lid (purple) completely encloses the drug. There's no way in or out when the lid is closed. So the lid has to open to allow drug entry/egress. Jeffrey created a nifty algorithm using a new method called QBITS, AF3, and LASErMPNN to design metastable lids that open w/o ligand
Ben Fry retweeted
We have a new paper out on bioRxiv describing how to turn "static" small-molecule binders into dynamic shape changers! Short 🧵
biorxiv.org/content/10.64898…

Ben Fry retweeted
Zero-shot design of drug-binding proteins via neural iterative selection−expansion
1. The work introduces NISE (neural iterative selection–expansion), a closed-loop algorithm that jointly optimizes protein sequence, protein structure, and ligand conformation—enabling true zero-shot design of small-molecule binders where prior deep-learning approaches struggled.
2. NISE alternates between two reciprocal neural conditionals: LASErMPNN samples sequences conditioned on a protein–ligand co-structure, and a co-structure predictor (RoseTTAFold-All Atom or Boltz-2) predicts the 3D protein–ligand complex from sequence + ligand identity; designs are selected by tripartite self-consistency (backbone and ligand r.m.s.d.) plus ligand confidence.
3. A key conceptual shift is treating “self-consistency” as protein–ligand self-consistency (not just foldability): many sequences can be backbone self-consistent, but far fewer place the ligand in the intended orientation; ligand self-consistency becomes a stringent computational screen for productive binding.
4. LASErMPNN is a ligand-aware heterograph message-passing model trained on PDB co-crystal structures to decode both amino-acid identity and side-chain dihedrals; it includes a pretrained ligand encoder trained on quantum-derived atom properties (for example partial charges), improving generalization to new ligands and reducing design pathologies like overpacking.
5. On exatecan (a camptothecin payload with a hydrolysis-prone lactone), NISE redesigned a four-helix bundle binder from scratch starting only from backbone + docked ligand coordinates; all 4 experimentally tested NISE designs bound exatecan (100% hit rate), with the best binder EPIC at Kd = 120 nM.
6. Against a traditional COMBS + Rosetta pipeline (16 tested designs), only 3 bound exatecan and the best was ~70-fold weaker than EPIC; analysis suggests NISE’s advantage comes from iterative “resculpting” of backbone/packing and ligand placement, not just filtering with a structure predictor.
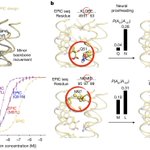
7. The authors then perform purely in silico affinity maturation (“neural proofreading”): LASErMPNN proposes single-site substitutions that reduce sequence NLL in the bound context; two substitutions (Q51N and M97L) each improve affinity >10×, and the double mutant improves EPIC ~100× to Kd = 1.2 nM without experimental feedback.
8. X-ray structures of EPIC (2.0 Å) and EPIC(Q51N) (2.2 Å) validate the designed binding mode and explain the affinity gain: Asn51 enables deeper burial and bidentate H-bonding to the hydroxyl and lactone carbonyl, with only ~sub-Å deviations from intended placement in the pocket core.
9. Functionally, EPIC variants protect exatecan’s lactone from hydrolysis: EPIC(Q51N/M97L) keeps >99% of exatecan in the ring-closed (bioactive) form for at least 50 hours in PBS pH 7.4, and protection persists even with high human serum albumin present—supporting drug-stabilizing delivery/sponge applications.
10. Generality is shown on apixaban using NTF2 scaffolds and Boltz-2 within NISE: 5 of 6 tested designs bound tightly (83% success), with the best (APEX) reaching Kd = 80 pM—nearly 10,000-fold tighter than prior LigandMPNN/Rosetta apixaban binder design reports on the same backbone family, and approaching the native target factor Xa affinity.
💻Code: github.com/polizzilab/LASErM… ; github.com/polizzilab/NISE ; github.com/benf549/CARPdock
📜Paper: doi.org/10.1038/s41586-026-1…
#ProteinDesign #ComputationalBiology #DeepLearning #StructuralBiology #DrugDiscovery #GenerativeModels #ProteinLigand #DeNovoDesign #GNN #Boltz2 #RoseTTAFoldAllAtom

Ben Fry retweeted
Great to see @benf549's and Kaia Slaw's work out in @Nature! Short thread about the work here: bsky.app/profile/nickpolizzi…
Ben Fry retweeted

Closing the design loop: two coupled neural networks for zero-shot small-molecule binder design
Protein structure prediction has gotten very good, but design is a different problem. Predicting how a known sequence folds is one thing, inventing a brand-new protein that grips a specific small molecule tightly is much harder, because you have to optimize three coupled things at once: the amino-acid sequence, the backbone it folds into, and the conformation of the ligand in the pocket. Change one and the other two shift. Deep-learning methods that crushed folding have mostly stumbled here, which is why most successful drug-binding proteins still come from slow experimental screening.
Benjamin Fry and coauthors close this loop with NISE (neural iterative selection-expansion). The idea is to pair two neural networks that solve reciprocal halves of the problem and let them refine each other. One, a graph neural network they call LASErMPNN, proposes many candidate sequences for a given backbone and docked ligand. The other, a structure predictor like RFAA or Boltz-2, folds each candidate and predicts where the ligand actually ends up. Designs that agree with themselves across both networks (low r.m.s.d. on backbone and ligand) get selected and expanded in the next round. No physics-based energy function is involved. The system climbs toward a high-probability mode of the joint sequence-structure-ligand distribution learned from the Protein Data Bank. When they swapped the neural predictor for a Rosetta energy function, the loop simply failed to optimize.
The results are hard to argue with. On the anticancer drug exatecan, all four tested designs bound it, the tightest about 70-fold better than the leading prior method. For the anticoagulant apixaban, NISE hit an 83% success rate using the same starting scaffolds a previous method had used, and its best binder reached 80 pM affinity, roughly 10,000-fold tighter and rivaling its natural target at a third the size. The same network then "proofread" its own design, suggesting two mutations that improved affinity 100-fold with no experimental input.
The practical signal is that bespoke small-molecule binders are becoming a design problem rather than a screening problem, and a fast one: a typical run takes about five hours on four GPUs. In drug delivery, biosensing, antidote development, and diagnostics, that means custom proteins to sequester, stabilize, or detect a specific compound can be generated on demand and matured in silico, compressing timelines that once needed large wet-lab campaigns.
Paper: Fry et al., Nature (2026), CC BY 4.0 | doi.org/10.1038/s41586-026-1…

Ben Fry retweeted

Zero-shot design of drug-binding proteins via neural iterative selection−expansion nature.com/articles/s41586-0…
A nice (or should I say NISE?) article written about our work by Derek Lowe!
Looking forward to sharing results on the diverse compounds we've applied our method to bind that didn't make it into this paper, but will be published in the coming months.
science.org/content/blog-pos…

Ben Fry retweeted

Nature research paper: Zero-shot design of drug-binding proteins via neural iterative selection−expansion
go.nature.com/4eFKHHY
Ben Fry retweeted

I build little translation to Polish, awesome papper

Dwie sieci neuronowe, pracując naprzemiennie, zaprojektowały od zera białka wiążące leki z 100-krotnie wyższą celnością niż dotychczas. 🧬
Zamiast losowej ewolucji w laboratorium, algorytm iteracyjnie selekcjonuje i rozbudowuje struktury białkowe. W testach nowe białka wychwytywały małe cząsteczki leków z dokładnością porównywalną do naturalnych receptorów.
Otwiera to drogę do projektowania białek do precyzyjnego dostarczania leków w organizmie oraz neutralizacji toksyn.
#AI #biotechnologia


(9/10) All of the LASErMPNN & NISE code is open-source under an MIT license. Shoutout to Kaia Slaw my co-author and the awesome RA who did all of the experimental validation for this work. Check out my personal website for an interactive visualization of the binders we designed.
(10/10) Links Below:
Link to Paper📜 (Open Access): nature.com/articles/s41586-0…
Link to Code 🤖 (MIT License): github.com/polizzilab/NISE
Personal Website 🧔♂️ (Interactive Visualization): benf549.github.io/