

Official Twitter of the Salzman research group at Stanford Biomedical Data Science
Stanford, CA
Joined June 2017
- Tweets58
- Following22
- Followers302
- Likes14


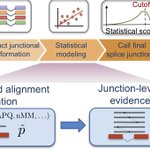



presentation by @TBaharav today at @RECOMBconf on the statistics behind SPLASH, a method that can be used as a robust alternative to one of the most common statistical tests
backlogged on posting our new work in genomics! using RNA splicing as a special case of expansive, ultra-efficient and fully statistical analysis of RNA or DNA sequencing data. cell.com/cell/fulltext/S0092…
Salzman Lab retweeted

Thrilled to share our new publication in PNAS on OASIS, an alternative to Pearson’s X² for analyzing contingency tables.
Made it to the front page! 1/ 7

Thrilled to announce 3 new papers from our lab, up on bioRxiv this past week.
biorxiv.org/content/10.1101/…
biorxiv.org/content/10.1101/…
biorxiv.org/content/10.1101/…

2. Design of a novel statistical test on contingency tables, OASIS, which provides interpretable and finite-sample valid rejection of the null, enabling reference-genome and metadata free strain detection in biological applications. Led by @TBaharav: biorxiv.org/content/10.1101/…

3. Development of NOMAD2, which enables reference-free biological discovery at unmatched scale and speed. In collaboration with @sdeorowicz and @marekkoki, led by @marekkoki and @roozbehdn, this package is publicly available on github: github.com/refresh-bio/R-NOM…

Salzman Lab retweeted
I can confirm that the NOMAD analysis method and the accompanying software is a game-changer for sequence analysis and discovery. Check this paper out! biorxiv.org/content/10.1101/…

Salzman Lab retweeted

NOMAD2 is an unsupervised, reference-free, ultra-fast next generation of NOMAD, revealing new insights into cancer transcriptomes. Joint work with @salzmanlab. Many thanks to all for great collab!
biorxiv.org/content/10.1101/…
Excited to share that our paper on reference-free transcriptomic analysis is live. biorxiv.org/content/10.1101/…

NOMAD+ discovers broad and new examples of transcript diversification in single cells, bypassing genome alignment and without requiring cell type metadata.
Salzman Lab retweeted
Had a blast at #biodata22! Was great to present and get feedback on NOMAD, our recent work for reference-free inference biorxiv.org/content/10.1101/….

Salzman Lab retweeted

"NOMAD is a single reference-free ‘omic algorithm for biological discovery across the tree of life"
Julia Salzman @SalzmanLab en #FrontiersInGenomics HOY martes 15 nov 5pm #EnVivo #EnLínea lcg.unam.mx/frontiers/live @ccg_unam @ibt_unam @unammorelos

Very excited that ReadZS, a scRNA-seq pipeline for detection of RNA processing, is out in @GenomeBiology! This work was led by @ea_meyer, K Chaung, and @roozbehdn.

ReadZS, from @ea_meyer, Chaung, @roozbehdn & Salzman, for identifying RNA processing (splicing, alternative polyadenylation etc) in RNA-seq data, without the need for pre-existing annotations. It works on both 3'-enriched and non-enriched data genomebiology.biomedcentral.…

ALT Schematic of how the method counts reads from each cell into windows
Excited to share our new preprint NOMAD! We present a unifying statistical formulation for many fundamental problems in genome science and develop an efficient reference-free algorithm (NOMAD) to solve it. Work by Kaitlin Chaung, @TBaharav, and @RZheludev. biorxiv.org/content/10.1101/…
Excited to announce that a project spearheaded between our lab and David Tse's has been selected for a @czbiohub Data Insights Award! Follow the link for more information.
chanzuckerberg.com/science/p…

Excited to share DIVE is live on bioarxiv! DIVE is a novel reference-free approach for de novo discovery of diversity-generating and mobile genetic elements implemented as a publicly available python package (work led by @jordiatjhu).

DIVE: a reference-free statistical approach to diversity-generating and mobile genetic element discovery biorxiv.org/cgi/content/shor… #bioRxiv
Very excited that our paper on the SICILIAN pipeline for robustly identifying splice junctions from scRNA-Seq led by @roozbehdn and @julia_eve_olivieri_ is out in @GenomeBiology! genomebiology.biomedcentral.…

SICILIAN can be applied to any type of scRNA-Seq (and even bulk RNA-Seq!) data. Thanks to our collaborators in @czbiohub, we have already applied SICILIAN to Tabula Microcebus and Tabula Sapiens, leading to organism-wide single-cell splicing analysis in human and mouse lemur
Finally, thrilled to announce that SICILIAN is available in both @nextflowio and @genomicscloud led by @olgabot and Ana Damljanovic, supporting the scale and precision needed for splicing analysis in massive scRNA-Seq datasets. (links available in github.com/salzmanlab/SICILI…)